Research Overview
At the ALS Hope Foundation, we strongly believe that basic research is a key tactic in our battle against ALS. Always pursuing innovative and collaborative projects, we are proud to support the work of the Dr. Robert Sinnott Research Lab at Temple University Lewis Katz School of Medicine.
Our Research Lab
The research lab is actively pursuing ways to better understand ALS and to treat and modify disease progression. The laboratory is small, so collaborations with other researchers and with industry are very important. Most of the work uses the SOD1 mouse model of ALS, which has been genetically engineered to carry the mutated human gene for SOD1 (superoxide dismutase). The mutant SOD1 gene is responsible for 20% of the cases of hereditary ALS in humans. The transgenic mice in the lab develop weakness and pathology that is very similar to people with ALS, providing an excellent model with which to study the disease. All supporters of the ALS Hope Foundation are invited to visit the lab and get a first hand glimpse of our research efforts.
Basic Research: Ongoing Projects
Genetic Modifiers of Disease in the SOD-1 and Dynactin Mouse Models
We have observed that survival in the transgenetic mouse model of ALS (G93A SOD1) is dependent on the mouse background. The original mouse model was created on a mixed background of two different strains called SJL and B6. When mice are mated to create either a pure B6 or SJL background, we observed that mice with a B6 background survive longer than those with the SJL background. We hypothesized that this was due to specific differences in genetic information between the two backgrounds, and that these genetic differences modified the severity of disease. In fact, we have now identified an area on mouse chromosome 17 that modifies disease, and our lab is actively working to identify the specific gene on this chromosome that is responsible in collaboration with Greg Cox at Jackson Laboratories. This project could provide a therapeutic target in human disease to help improve the outlook for people living with ALS. We are also studying whether this effect is observed for other models of motor neuron damage. In fact, we have now found similar differences in the severity of disease in the dynactin mouse, another genetically-engineered model based on the mutated dynactin gene, and we chromosome 17 modifiers may also be responsible in this case. This is exciting because it means that similar modifiers could affect many different types and models of motor nerve damage, so a possible treatment based on this information could be more universally applied. We will now be examining our potential modifying genes in other disease models including worm models. This work was recently published in the journal PlosOne.
Developing a Better Model of Human Disease: The VLE SOD1 Model
The mouse model of ALS most commonly used for experiments has 24 copies of the abnormal SOD1 gene; this mouse dies very young. Once the disease starts, there is a rapid progression. Since ALS in people occurs later in life and is slower in progression, we have developed a model of disease that starts later and progresses more slowly by decreasing the number of mutant SOD1 gene copies to only four. We call this our VLE (very low expression) SOD1 model and feel that it more accurately reflects the disease in humans. This work was recently published in the journal PlosOne. We have characterized this animal and are now using it to study early events and environmental conditions that may precipitate disease.
Developing Other Models of Motor Neuron Disease: Spastin Models
Hereditary Spastic Paraplegia (HSP) is a neurodegenerative disorder affecting only the upper motor neuron, resulting in motor system involvement similar to ALS and PLS. Mutations of the spastin gene are the most common cause, accounting for over 40% of HSP cases. We have now created a new genetic mouse with inducible expression of a human spastin mutantation andare characterizing the mouse to see if it exhibits motor weakness and disease. If the animal develops spasticity, it will provide a model of PLS and HSP to study mechanisms of upper motor neuron damage. This mouse will also be useful for design of treatment strategies for PLS and HSP as well as ALS.
Developing New Therapeutics
We are working with LifeSplice, a biotech company, to develop agents that can alter the form of proteins that are damaging to nerve cells. We call these agents splice modulating oligomers because they alter the way DNA is cut up or spliced to make RNA, the process that determines which variation of a protein is made by your cells. In people with ALS and other motor neuron diseases, these toxic proteins lead to nerve cell death. By altering these proteins, we might be able to slow or stop the disease process. Our splice modulating molecules are now being tested in animals for safety.
In addition, we are working in collaboration with the Pharmacology Department at Drexel University College of Medicine to test potential drugs that alter sigma 1 action on nerve cells in our mouse models of ALS. Sigma 1 signalling is important for nerve cell survival and function. A mutation in the gene for sigma 1 is responsible for some cases of familial ALS.
The MDA/ALS Center of Hope at Temple University Lewis Katz School of Medicine provides an important setting for research towards understanding, treating, and curing ALS. That is why the ALS Hope Foundation supports an extensive clinical research program at the Center of Hope. Without research, there would be no progress in our fight against ALS. Clinical Research is a broad term, encompassing clinical trials (studies on the effect of a medicine on the disease or its symptoms) and clinical research that examines other aspects of the illness, such as assistive technology or quality of life. Clinical research at the Center complements our basic research program, in the hope that promising therapeutics identified at the lab bench can be tested for efficacy in the clinic.
For more information on the clinical trials being conducted at Temple's ALS clinic, please contact our Research Nurse Coordinator, Kathleen Hatala, via email at Kathleen.Hatala@tuhs.temple.edu.
To learn more about each of these trials, please go to clinical trials.gov and search for ALS trials or go to the Northeast ALS Consortium site neals.org.
Clinical Research: Ongoing Projects
The Healey ALS Platform Trial
This trial is a novel design where multiple drugs will be tested in tandem against one placebo group. If volunteers with ALS meet a universal set of criteria, they are randomized into one of the treatments. The criteria are broader than we have had for most trials and allow wider access for PALS and the chances of treatment are higher than traditional trials as the placebo group is shared between them. The first 5 drugs were 1) Zilucoplan (UCB); 2) Verdiperstat (Biohaven Pharmaceuticals); 3) CNM-Au8 (Clene Nanomedicine, Inc); 4) Pripodine (Prilenia Therapeutics); and 5) Trehalose (Seelos Therapeutics). The 6th drug, 6) ABBV-CLS-7262 (Calico), targets eIF2B, a key regulator of the integrated stress response; 7) DNL343 (Denali), DNL343 is an investigational CNS-penetrant small molecule integrated stress response (ISR) inhibitor designed to activate the eukaryotic initiation factor 2B (eIF2B) and suppress aberrant ISR activation. Enrollment is currently completed for the 6th and 7th drug regimens.
ROAR DiGAP
Several years ago, in partnership with patients, Dr. Richard Bedlack of Duke University started a novel program called ROAR (Replication of ALS Reversals). This consists of small pilot trials of vitamins and supplements that are associated with cases of recovery from ALS (“ALS Reversals”). Unlike more traditional trials, ROAR trials are widely inclusive and have historical rather than placebo controls (all enrolled patients get the treatment under study at no cost to them). While this program has not yet found a treatment that reverses ALS, its trials enroll and get answers quickly and at a fraction of the cost of most other trials.
We are thrilled to be partnering with Dr. Bedlack on his next ROAR trial. This will utilize a new technology called DiGAP (deep integrated genomics analysis platform) to determine which of 4 biochemical pathways is most likely killing motor neurons in each enrolled patient. Based on this, each patient will receive a personalized treatment tailored to their own biology. Each of the 4 treatments being offered is associated with at least one ALS Reversal. Enrollment is complete for this trial.
In addition, the clinic is actively helping to find new ways to diagnose disease, track the progression, and study the response to treatment. The following studies are open to enrollment at the MDA/ALS Center of Hope.
Oxidative Stress Response in Different ALS Types
In a recent study, Radicava (edaravone) has demonstrated slowing of disease progression in ALS. It has been shown to inhibit motor neuron death in animal models by reducing oxidative stress. Given that oxidative stress plays a role in ALS; there is evidence that Radicava can affect disease progression by reducing oxidative stress. Biomarkers of oxidative stress and antioxidant capacity can be measured in body fluids (plasma, cerebrospinal fluid (CSF), and urine), thus the effect of Radicava on oxidative stress can be evaluated in a systematic way by comparing levels before and after treatment over time and compared to changes in an untreated population of PALS. The purpose of this study is to determine if different types of motor neuron involvement in PALS vary in baseline levels of oxidative stress and whether any subgroup has a selective benefit as measured by a series of oxidative markers.
Natural History of ALS
As part of the ALS Natural History Consortium we are collecting clinical data on people with ALS and other motor neuron diseases that attend the Center of Hope and entering the de-identified information into a large multicenter database called Neurobank. This data will allow comprehensive characterization of the clinical picture of ALS and allow us to study the disease and characteristics that will predict outcomes, stratify populations, as well enable clinical studies. By using a global identifier for each clinical record, the clinical data can be tied to clinical samples.
PLS Natural History Study
The goal of the PLS Natural History study is to advance the current research state of PLS by generating collecting clinical data and biological samples over time to understand the natural progression of the disease. This will advance development of future research and clinical trials. If you consent to participate in this study, you will have study visits every 3 months for 1 year and a final study visit at 2 years (24 months). Visits 2 (3 month follow up), 4 (month 9) and 6 (month 24) will occur over the phone and visits 3 (month 6) and 5 (month 12) will be in-person. In addition to collection of clinical data relevant to PLS an electromyography (EMG) test will be performed at visit 5 (month 12).
MDA MOVR Data Hub
The Muscular Dystrophy Association (MDA) wants to collect information about people with neuromuscular diseases to better understand their natural disease progression and ultimately improve their medical care, quality of life and survival. If you consent to participate in this study, you will be asked to provide clinical information from your medical chart so long as you are being followed at our clinic (you can withdrawal at any time). The information collected from your medical chart and provided by you will be stored in a protected database called the MDA MOVR Data Hub.
Tissue Bank
In a disease like ALS in which the cause is unknown, no animal models of ALS can substitute for understanding how it affects humans. We are currently collecting blood, urine, CSF and autopsy materials from people with ALS and other motor neuron diseases to look for clues in the human tissue. All samples are tied to de-identified clinical information in a database to help maximize the usefulness of this precious resource. This includes demographic information; environmental exposures; and medical history. This will increase availability of human tissue for research with pertinent corresponding clinical information.
Predictors of Respiratory Events in ALS
The goal of the PALS Cohort study is to understand if certain characteristics of ALS specific to you may help predict the risk of developing problems with breathing and whether they determine how well breathing function will be over time. The study will be enrolling people who have been diagnosed with ALS in the past 5 months prior to enrolling in the study. If you consent to participate in this study, clinical data relevant to your diagnosis, vital signs, breathings test results and use of non-invasive ventilation or NIV (breathing equipment) and tracheostomy placement will be collected at 5 clinic visits over about 12 months. A secondary goal, through an optional sub-study, is to gather patient concerns, expectations, and knowledge regarding early respiratory care in ALS. This includes an approximately 30 minutes recorded, transcribed and de-identified phone call.
Evaluation of GI Symptoms in ALS
The gastrointestinal (GI) tract has a wide array of symptoms that has been investigated in ALS. This may include gastric motility and delayed emptying indicating autonomic dysfunction that may occur in patients with ALS. Some of the typical symptoms frequently experienced include constipation and bloating and may be attributed to decreased physical activity, dysphagia resulting in inadequate fiber intake, or dehydration. It is important to adequately determine the frequency by which these symptoms are experienced by ALS patients. Evaluation of symptoms will consist of completing a questionnaire.
The following are projects aimed at evaluating the effectiveness of some of these new technology developments for communication and control for individuals with ALS.
EEG-Based Brain-Computer Interface
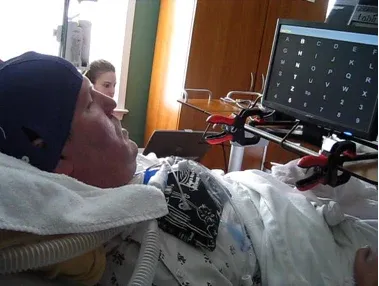
By using EEG (brain wave) signals from the scalp to create a signal, the Brain-Computer Interface (BCI) allows people to make selections from the computer screen. The study is intended to evaluate both the complexity of the system and the degree to which each participant will be able to communicate. Trials will consist of asking the subject to follow a series of simple instructions and to complete certain tasks while using the BCI. This study is open for enrollment.
Assessing Assistive Technology Use and Needs
The purpose of this study is to assess the current assistive technology use and needs of people living with amyotrophic lateral sclerosis (ALS) and other neuromuscular diseases. This data will be used to determine the level of familiarity and effectiveness of this technology. Our hypothesis is that there are 1) gaps in the information that people with neuromuscular diagnosis have about assistive technology and that there are 2) areas of need for AT that are not being addressed by designers. This information gathered can then be used to 1) develop educational resources to provide information to individuals who need the AT and 2) provide a resource to individuals developing AT to address the needs of their target population. This study is open for enrollment.
Key Terms
- Biomarkers
- Distinct biochemical, genetic or molecular characteristic that is an indicator of a disease
- PALS
- Person living with ALS Placebo: A control substance given with no therapeutic or treatment effects expected
- Placebo-controlled
- Trials with two groups of people, those who receive the medication or treatment being tested and those who receive a placebo treatment.
- Randomized
- Participants are selected to be placed by chance, like in the flip of a coin, to either the placebo or active treatment group